
protocol overview
Induction temperature and inclusion bodies
Inclusion bodies are insoluble aggregates from overexpression. A cooler induction can leave more protein soluble, and a gel of soluble and pellet fractions
- Author
- EVRINTH Editorial Team
- Published
- 8 October 2026
- Updated
- 8 October 2026
- Reading time
- 10 min
A band that appears only after you spin a lysate hard is a different result from a band in the clear supernatant. Inclusion bodies are insoluble aggregates of protein made faster than the cell folds it, and the induction temperature is one of the few variables a cloning laboratory can change without redesigning the gene. The decision this overview supports is whether a cooler shift, of the kind the vector protocol allows, moves your research protein into the soluble fraction, judged on a gel. The host is a non-pathogenic E. coli expression strain already accepted by your institution. How that host is grown and transformed is bacterial culture and transformation. How to look at the lanes afterwards is reading a protein gel.
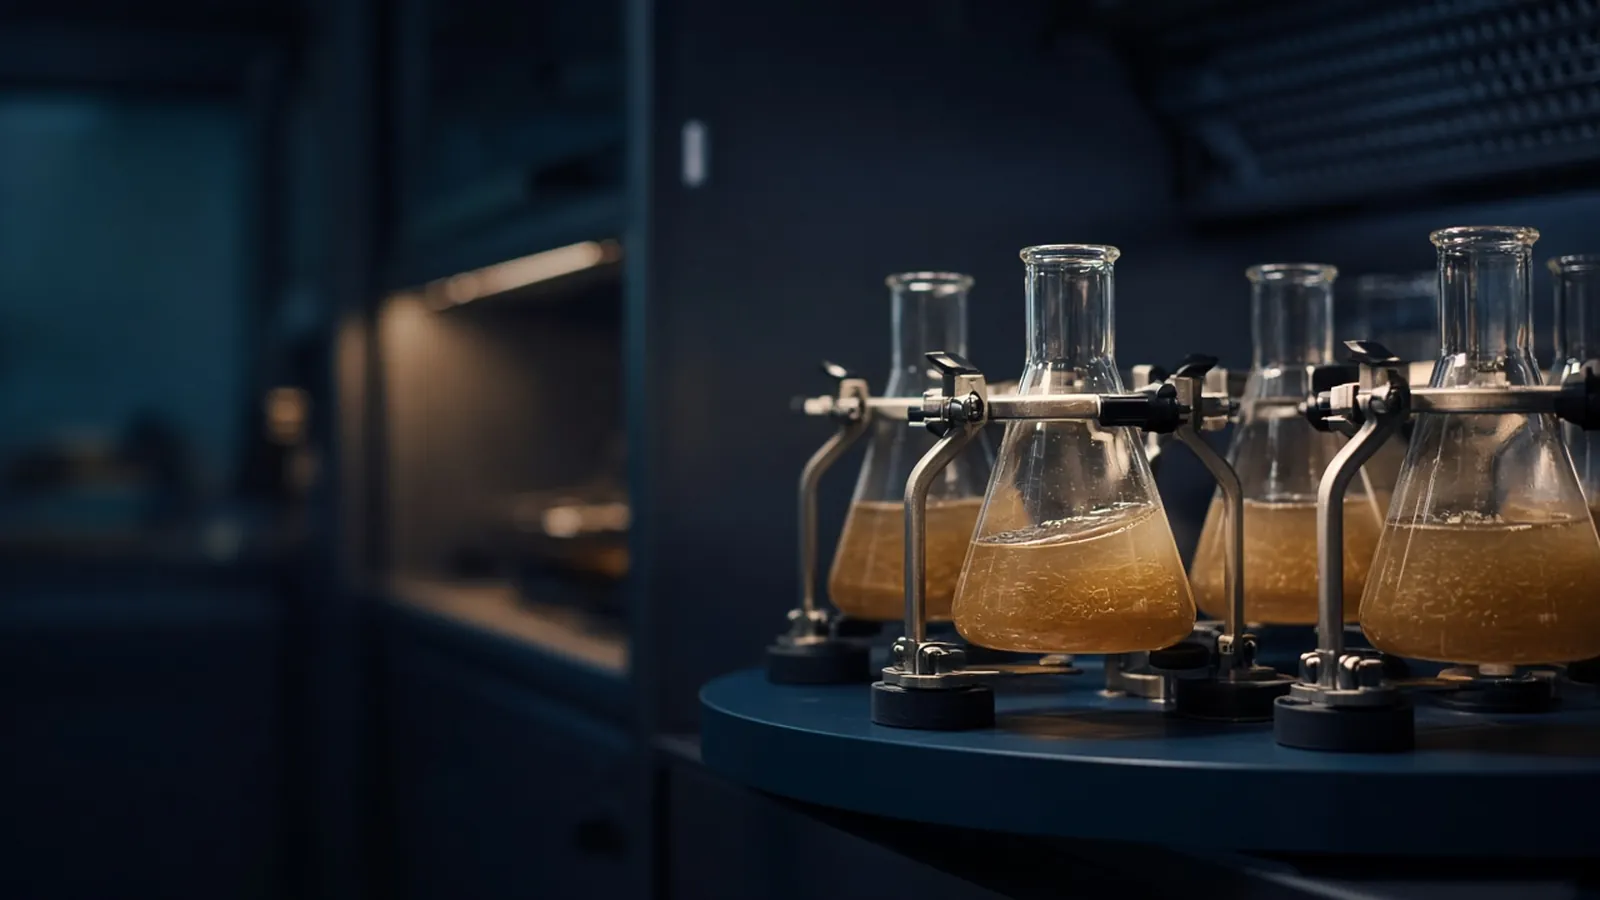
The hero photograph shows Erlenmeyer flasks of amber culture on an orbital shaker. Those flasks are the induction vessel. They are not a chromatography column, and this page does not become one. Medium, inducer and antibiotic classes are in the reagents and chemicals catalogue. The wider path is the molecular biology pathway. Sequence design, including codon choice that sometimes affects expression, can be framed from custom gene synthesis as an enquiry reference only. A specification goes through the quote request.
What the pellet actually is
After you break the cells and centrifuge, the supernatant holds what stayed soluble. The pellet holds unbroken cells, membranes, and, when overexpression has been fierce, a dense mass of the recombinant polypeptide. That mass is the inclusion body. On a stained gel it is often the dominant band in the insoluble lane and a faint or missing band in the soluble lane. The stain does not prove identity. A specific blot, an activity assay, or a mass result does that later. The gel's job at this stage is the split between fractions.
Strong bacteriophage-T7-style systems in DE3 lysogens, and other high-level promoters used in K-12 expression hosts, are good at filling that pellet. The cell is not failing so much as obeying a promoter that outruns folding. Lowering temperature after induction slows translation and growth. Chaperones and the chain itself then have a better chance with some proteins. The shift people describe is commonly toward the mid-20s Celsius, and some vector sheets go lower still, into the high teens. Follow the vector protocol. A number remembered from a different promoter is not that protocol.
It is not magic. A protein with several disulphide bonds is in the wrong cellular compartment for those bonds in the E. coli cytoplasm, cold or warm, unless the strain was specifically built to change that redox environment and your protocol says to use it. A membrane protein may remain a pellet for reasons that temperature will not touch. A polypeptide that needs a modification the bacterium does not perform can be soluble and still inactive. Say that before you spend a week shifting flasks. Background on how cells handle proteins is in the NCBI Bookshelf text of Molecular Biology of the Cell. A public record of a named protein, if you need the real sequence rather than a bench nickname, can be checked in UniProt.
Host, vector, inducer, antibiotic
The host has to match the promoter. A DE3 lysogen is there so a T7 promoter has a polymerase to call. A plain cloning strain with the same plasmid will look like "no expression" at every temperature. The vector sheet names the inducer. For many lac-operator systems that inducer is IPTG, used in a concentration class the sheet prints, commonly from sub-millimolar to about one millimolar. Autoinduction media, where lactose metabolism leads to induction without a timed IPTG addition, are a different class. Do not mix the two stories in one flask.
Antibiotic selection stays in the preculture and, unless the sheet says otherwise, in the expression culture. Plasmid loss is fastest when the protein is a burden. A cooler flask does not reintroduce a plasmid the cells already dropped. Inoculate from a single colony on a fresh selective plate. The practical shape of starting that culture is the Addgene protocol for inoculating a bacterial culture. Enzyme and buffer classes around a later analysis sit among Promega protocols. They are not a lysis recipe for this page.
Density at induction is a window, not a vibe. Many T7 protocols name a mid-log turbidity at 600 nanometres, often printed somewhere around 0.4 to 0.8 on the spectrophotometer that laboratory calibrated. Your vector sheet wins if it names a different window. Turbidity is not a cell count until you have related the two. It is good enough to stop you inducing a culture that is already stationary.
The comparison worth running
Grow two flasks, or split one culture, so the only deliberate change is temperature after the inducer goes in. One stays at the growth temperature. One moves to the cooler set point the protocol allows, in a shaker that can hold it, with the clamps tight and the fill low enough that the cooled culture still gets air. A cold flask with no agitation becomes a hypoxic stationary culture and confounds the temperature story. Take an uninduced sample. Take a sample at the harvest time the protocol names. Do not invent a 16-hour default if the sheet says four.
Lyse by the method your laboratory already validated for this host. Separate a clarified soluble fraction from the pellet. Load amounts you can compare, with a marker, and read the gel as the protein-gel article describes. Then branch. Soluble band only under the cool shift: keep that condition and stop calling the protein absent. Pellet under both: temperature was not the lever, or not the only lever. The next change might be inducer concentration class, a different host the committee has already approved, or a sequence change discussed as a design question. Both fractions: you may work with the soluble material and treat the pellet as a separate, later question. No band anywhere, including a total-lysate lane: suspect the strain, the inducer, the antibiotic, or a frame error, before you buy a colder shaker.
This overview stops at that split. It does not set lysis-buffer recipes, refolding steps, or column methods. Those are a different method, with their own controls, and they are not required to answer the temperature question.
How to read the three outcomes
| Where the candidate band sits | What you can claim | What you do next |
|---|---|---|
| Mostly in the soluble fraction | Under this induction, a useful share stayed soluble. Identity still needs a specific test. | Keep the condition. Confirm the band before you scale the flask. |
| Mostly in the pellet | Most of the stained product is insoluble. A cooler shift is one variable, already tested or still available. | If you have not shifted, run the protocol's cooler arm. If you have, temperature alone did not solve it. |
| In both fractions | The culture split the product. The soluble part is the part you can see without a refolding story. | Harvest the soluble fraction for whatever the protocol allows. Do not pretend the pellet is the same sample. |
| Missing from both, uninduced also empty | Nothing on the gel matches an induced product. | Check promoter, host genotype, inducer, plasmid retention and the reading frame before another temperature. |
When the cooler flask changes nothing
A heat-only shaker whose panel says 18 Celsius in a room at 30 Celsius is not running the protocol. Measure the broth. A culture induced in late stationary phase is also not the protocol, and cooling it late will not unwind a pellet that formed for hours. An uninduced lane that already shows the band means the promoter is leaky. Temperature still matters, but you no longer have a true off state, and any toxicity of the product starts before you add inducer.
A band that appears in the pellet of a control strain with no insert is not your protein. Host proteins form pellets too when lysis is harsh or the spin is long. Comparable loading, an uninduced lane, and a specific identity test keep that mistake small. If the polypeptide itself is hazardous, a toxin or anything the committee has not assessed, stop before induction. A temperature shift is not a containment decision.
Safety and research use
Expression work uses the same cloning or expression strain the institution already assigned, at the volume they approved. Scaling from a shake flask to litres is a change you report. Antibiotics in the medium are chemicals. Waste culture is inactivated as the safety office requires. This page is not medical advice, not a purification, and not a method for producing a hazardous protein. If the insert changes the risk, the assessment changes before the shaker does.
A hot building and a set point that lies
The operational failure in a warm climate is a machine that only heats. You can grow at 37 Celsius easily, and you cannot shift toward the mid-20s unless the shaker refrigerates. Write "cooling required" into any equipment note, plus the temperature class and the orbit that keeps a half-empty flask aerated at the cooler speed you will use. A power cut during the shift splits your comparison: the flask that warmed back to ambient is not the cool arm. Discard that pair and repeat. Humidity on the shaker lid drips into loose closures. Use the closure the flask method already specifies, and label the real temperature you measured beside the panel value.
What to send when you ask about an expression host or a shaker
Name the strain class, the promoter class, the antibiotic, whether you need a cooler induction, and the temperature the protocol actually states. Ask whether a shaking incubator cools or only heats, and ask for the orbit class and the clamp style without naming a favourite brand. The expression method can be discussed against the vector sheet. Identity of the polypeptide, if the sequence is still open, stays a question on the custom gene synthesis reference. Send the practical fields through the quote request, and keep the gel plan attached so soluble and pellet are both part of the work you describe.
Compare a warm induction with a cooler shift before you call the protein absent
- 01Grow at the strain's normal temperatureStart from a single colony on the correct antibiotic and grow the expression host at the temperature that strain uses for ordinary growth, often near 37 Celsius for routine E. coli. Follow the vector protocol for the density window at which you induce. Keep the flask aerated.
- 02Add the inducer the vector sheet namesUse the inducer class and the concentration class printed for that promoter. Many lac-controlled systems are induced with IPTG in a sub-millimolar to about one-millimolar class. The sheet rules. Save an uninduced sample from the same culture so the gel has a baseline.
- 03Shift temperature only if the protocol says toIf the vector or the protein method calls for a cooler induction, move the flask to a shaker that can actually hold that temperature, commonly a shift toward the mid-20s Celsius or lower. Keep shaking. A heat-only machine in a hot room will not perform a cold shift.
- 04Read soluble and pellet fractions on a gelLyse by the method your laboratory already uses, separate a clarified supernatant from the pellet, and load comparable amounts. How to read the lanes is in the protein-gel article. A strong band only in the pellet is an inclusion-body result. It is not a purification method, and it is not proof the band is the right protein.
Questions from the bench
Does a lower temperature guarantee soluble protein?
It slows growth and translation and often raises the soluble fraction of proteins that were merely being made too fast. It does not fold every polypeptide. Proteins that need disulphide bonds, membranes, or modifications E. coli does not supply can stay insoluble or inactive at any temperature the shaker can hold. The gel decides for this construct, not a general rule.
Why is the pellet not automatically a failed experiment?
An inclusion body is a concentrated form of the overexpressed polypeptide, and some research programmes deliberately use that pellet. This overview does not give a refolding or chromatography recipe. It tells you to see which fraction holds the band before you choose a path the vector protocol actually contains.
Should the antibiotic stay in the expression flask?
Keep the selective drug in the culture unless the vector protocol gives a specific, short reason to omit it and a way to check the plasmid is still there. Cells that drop a burdensome expression plasmid outgrow the cells that keep it. A gel with no band is sometimes a plasmid-loss result, which a lower temperature will not repair.
What if the shaker cannot cool below the room?
Then you cannot claim a mid-20s or a colder induction, whatever the set point says. Write the real temperature of a flask, not only the panel, into the notebook. In an enquiry for a shaking incubator, ask whether the machine cools or only heats, and name the temperature class the protocol requires.
References
Manufacturer names identify published method classes. Trademarks remain with their owners. Catalogue records on this site are independent references for enquiry. They are not a statement of inventory, distribution rights or a supply commitment. This page is educational. It is not medical advice, a diagnostic protocol or a biosafety approval.
Catalogue
Related products and categories
These links follow the subject of the article into published manufacturer references. A listing is a reference for an enquiry, not a statement of stock or distribution rights.
Continue in this cluster
Related reading
Bacterial culture and transformationHow non-pathogenic E. coli cloning strains are grown and how plasmid DNA is introduced, with the controls that make a colony meaningful. Reading a protein gelHow to read a protein gel: what SDS-PAGE bands, smears, ladders and loading differences can support, and what they cannot identify.A glossary of bench microbiology termsPlain definitions of colony, lawn, streak, pellet, transformant and the other bench words a cloning afternoon depends on.