
selection guide
His-tags and why a band is not pure protein
A His-tag elution can look like one band and still be a mixture. IMAC is a capture step, and a second method is what supports a purity claim.
- Author
- EVRINTH Editorial Team
- Published
- 8 October 2026
- Updated
- 8 October 2026
- Reading time
- 6 min
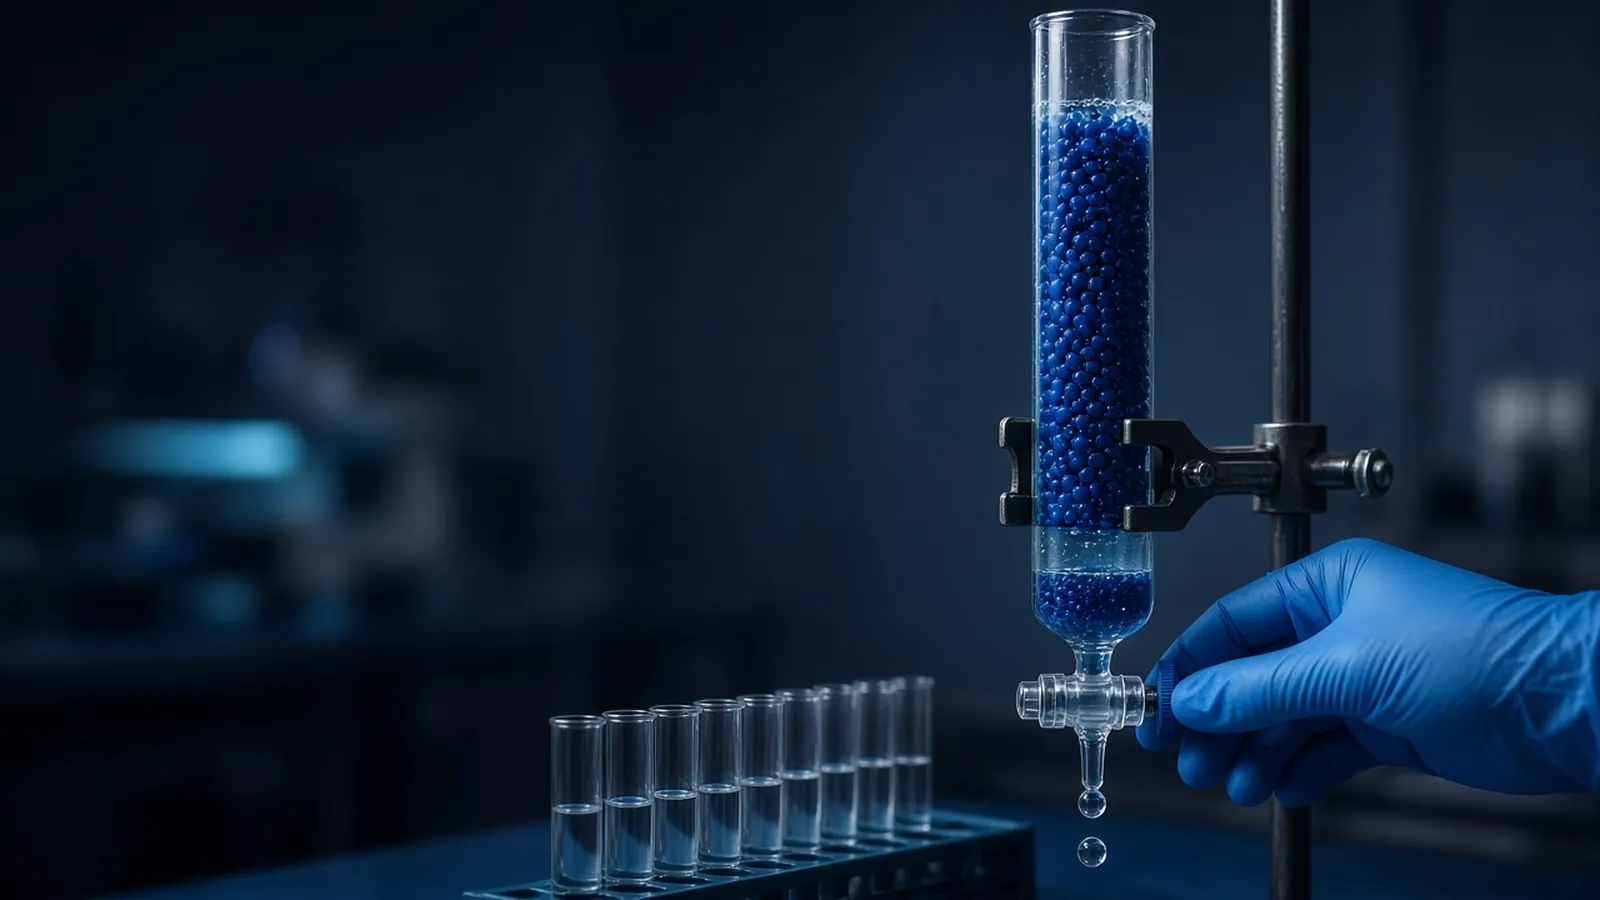
Histidine tags made metal-affinity capture ordinary, and they also made a single blue band look more trustworthy than it is. Immobilised metal affinity chromatography, usually on nickel or cobalt, binds clusters of histidines. Your tag is one such cluster. It is not the only one in a lysate. This selection guide is about what an elution band allows you to say, and when you still need a second step. The broader capture logic is in affinity and ion-exchange protein purification. How to look at the lane without renaming it as identity is in reading a protein gel.
What the resin is selecting
A nitrilotriacetic acid or similar chelator holds a metal ion, and the metal coordinates histidine side chains. Imidazole competes for those sites, which is why a low imidazole wash removes weak binders and a higher imidazole step elutes the tighter ones. Cobalt resins are often described as stricter than nickel resins. Stricter is not specific to your construct. It is a different balance of affinities. Neither metal reads the plasmid name.
Host proteins with natural histidine-rich stretches survive that balance often enough to be famous. In E. coli preparations, proteins such as SlyD are discussed again and again because they co-purify. You do not need to memorise the list to use the lesson. Expect extra bands. A wash strategy and a second chromatography step are how you respond. If the product was expressed as an inclusion body and then refolded, the His-tag capture still does not prove the refold worked. That comparison is in inclusion bodies and refolding as a concept.
A band the gel cannot split
SDS-PAGE separates mainly by polypeptide length. Two proteins of similar mass land on top of each other. One stained band can be a mixture the eye will never resolve, especially if one species dominates. Loading less can sharpen the picture. It cannot invent resolution the gel percentage does not have. An orthogonal method does: ion exchange, size exclusion if the shapes differ enough, a specific blot, activity, or mass spectrometry of the band. HUPO is a useful doorway into the proteomics side of identity. A single elution peak on a chromatogram has the same limit. Peak shape is not purity.
Quantifying that lane with a Bradford assay or A280 and reporting "95 percent pure" from a glance is a category error. Densitometry of a Coomassie gel can support a cautious statement about stained material at that load. It still misses what the stain does not see, and it still merges co-migrating bands. Say what you measured.
Buffers that quietly empty the column
Imidazole in the lysis buffer must be low enough that the tag still binds, and high enough in the wash to matter. The numbers belong to the resin protocol, not to a blog. EDTA and other chelators remove nickel and cobalt. A protease-inhibitor tablet tossed in because the box said "EDTA" can strip the column. The protein then flows through, and the gel of the elution is empty for a mechanical reason. Reducing agents are the other compatibility trap. Some resins accept a low millimolar level of DTT or of TCEP. Some do not. The difference is written on the resin product information. Follow that note. A column run with the wrong reductant can lose capacity or precipitate.
Clarify the lysate before it meets the resin. A cloudy load plugs the bed and smears the elution. That problem is physical, and it is shared with every other column.
Cleavage and the second step
A protease site between the tag and the protein is useful when the tag must leave. Thrombin, TEV protease and their relatives are choices with different scar sequences. The protease is now a contaminant. So is the cut tag, which may itself stick to the metal resin if you try a reverse-IMAC without thinking. Plan the polish: the protease is removed by its own tag, by ion exchange, or by size, according to a design you wrote down first. Incomplete cleavage leaves a doublet. Report both species or cut again. Do not average them into one molecular weight.
Choose a second step when the use of the protein cares about contaminants: enzymology, structural work, binding constants, or anything you will call pure in a methods sentence. For a quick activity screen of a well-behaved enzyme, a careful IMAC elution with documented extra bands may be enough if you say so. The selection is about the claim, not about a moral ranking of tags. The construct sequence, including the tag, has to match the mass on the gel and any extinction coefficient you apply.
| What you see | Consistent with | Still possible |
|---|---|---|
| One elution band | Strong enrichment at this load | A co-migrating host protein or a fragment |
| Several wash bands, one late band | A useful imidazole window | Overloading that hides faint contaminants |
| Empty elution | Tag missing, EDTA present, or protein insoluble | Expression failure, which a pellet gel would show |
| Doublet after cleavage | Partial tag removal | Proteolysis at a second site |
Failure modes in a sensible order
Flow-through that still contains the target means the tag is inaccessible, cleaved already, chelated away, or the resin is saturated or stripped. Check a gel of the flow-through before you induce another litre. A target that stays on the resin until harsh imidazole, beside a crowd of other bands, needs a better wash window or a different metal, then a polish. An elution that clouds as imidazole is diluted has an aggregation problem, not a tagging success. Clarify before the next column.
Protease inhibitor choice interacts here. EDTA protects against metalloproteases and destroys the IMAC step. Serine-protease inhibitors can block the tag-cleavage enzyme you add later. Those collisions are why the inhibitor list is part of the purification plan. Write the resin metal, the imidazole steps you actually used, and the load volume in the same note as the gel. A later repeat cannot reconstruct a wash you only remember as "a bit of imidazole".
Research limits and the bench
Nickel and cobalt salts, imidazole and the buffers around them are handled under ordinary chemical hygiene. This guide is not a therapeutic protein specification and not biosafety approval for the expression strain. Follow institutional rules for the host.
A pump that stops during a power cut leaves a column half-washed and warming up. Do not resume and call the elution equivalent to a controlled gradient. Note the stop, and consider whether the bed has dried. In a warm room, imidazole buffers and clarified lysates are not day-long bench ornaments. Keep the load cold if the protein needs it, and write the temperature you actually held.
What a sourcing enquiry should include
State the tag sequence, the host, whether the tag must be removed, which metal chemistry you have in mind, and what purity claim the downstream experiment needs. Resin class and buffer components are a reagents and chemicals conversation. Send that specification with a quote. If the question is really about an expression and purification plan, use the custom protein expression and purification page as an enquiry reference. Discuss the method there. Do not read it as a statement that a His-tag purification is already being performed.
Questions from the bench
Does one band in a His-tag elution mean the protein is pure?
It means the stained material in that lane migrates as one visible species at that load. A contaminant of similar mass can hide inside the band, and a faint contaminant can sit below the detection of a heavy load. Purity is a claim about other methods as well as about the gel.
Why do host proteins bind a nickel or cobalt column?
Immobilised metal affinity chromatography binds histidines, not a magical tag detector. Some host proteins carry natural histidine stretches and co-elute. A wash at a modest imidazole concentration removes a subset of them. It does not repeal the chemistry.
Can I add EDTA or DTT to every His-tag buffer?
Not safely as a habit. EDTA chelates the metal and can strip the resin. DTT and other reductants are tolerated differently by nickel and cobalt resins and by different manufacturers. Follow the resin note for the bottle you have. A stripped column looks like a failed expression.
When is tag cleavage worth doing?
When the tag changes the mass you must report, interferes with activity, or must be absent for the next experiment. Cleavage only helps if it goes near to completion and a second step removes the protease and the free tag. A gel of the digest is the check, not the supplier's theoretical cut site.
References
Manufacturer names identify published method classes. Trademarks remain with their owners. Catalogue records on this site are independent references for enquiry. They are not a statement of inventory, distribution rights or a supply commitment. This page is educational. It is not medical advice, a diagnostic protocol or a biosafety approval.
Catalogue
Related products and categories
These links follow the subject of the article into published manufacturer references. A listing is a reference for an enquiry, not a statement of stock or distribution rights.
Continue in this cluster
Related reading
 Reading a protein gelHow to read a protein gel: what SDS-PAGE bands, smears, ladders and loading differences can support, and what they cannot identify.Affinity and ion-exchange protein purificationHow affinity capture and ion exchange pull a protein from a lysate, and why each collected fraction still needs an identity check.Inclusion bodies and refolding as a conceptAn inclusion body is insoluble expression, not proof a protein can never fold. Refolding is empirical, and activity rather than a band shows success.